


Metabolic diseases occur when the body's biochemical machinery fails to function correctly. Under normal physiological conditions, metabolism requires a precise balance of substrates (such as carbohydrates, fats, and proteins), properly functioning enzymes, and sufficient levels of co-factors and co-enzymes. When an enzyme is deficient or mutated, the metabolic pathway is disrupted, often leading to the toxic accumulation of substrates or the deficiency of essential end-products.
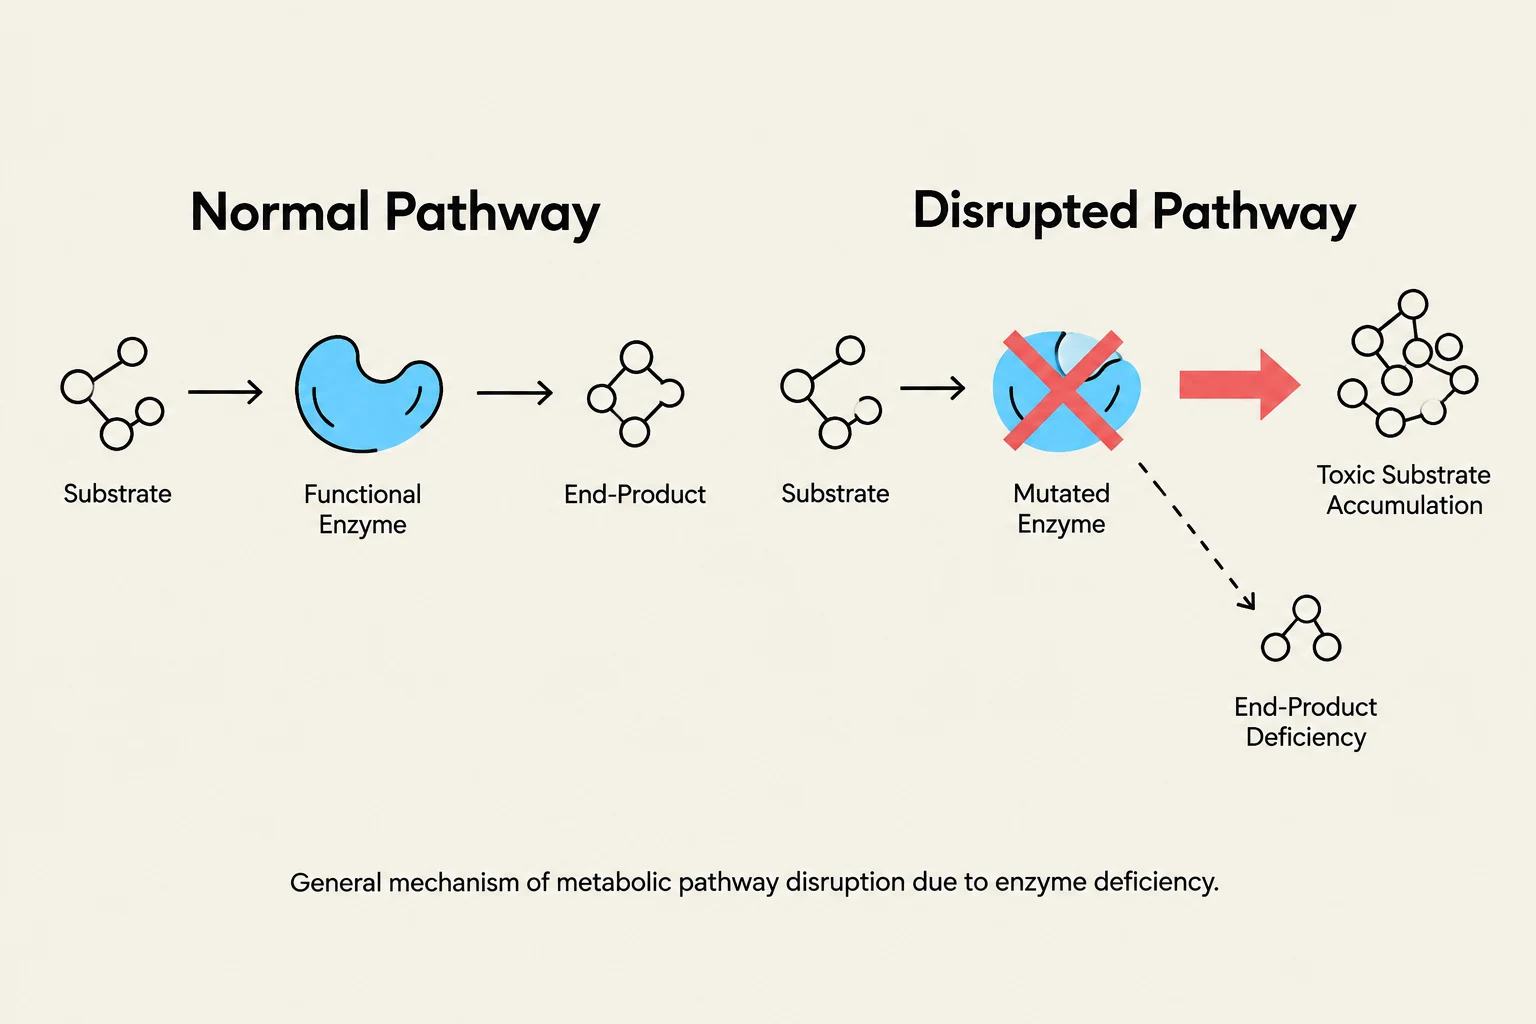
Two significant examples of such genetic, autosomal recessive disorders are Hereditary Fructose Intolerance (HFI) and Phenylketonuria (PKU). While both involve enzyme deficiencies, they impact different macronutrients and present distinct clinical challenges.
Hereditary Fructose Intolerance (HFI)
Hereditary Fructose Intolerance is an autosomal recessive disorder that specifically disrupts fructose metabolism. In a healthy individual, the liver processes various sugars to maintain energy homeostasis. However, individuals with HFI suffer from a deficiency in the enzyme Aldolase B.
The Role of Aldolase B and Fructose Metabolism
The primary issue in HFI is the inability of the liver to effectively break down fructose. It is important to note that this deficiency is highly specific to the Aldolase B enzyme. The body possesses other isoforms, namely Aldolase A and Aldolase C, which allow the individual to continue metabolizing glucose normally.
Despite the ability to process glucose, the disruption in fructose metabolism has a cascading effect on the body's overall metabolic state, specifically impacting glucose levels. When fructose cannot be processed, it can interfere with the liver's ability to maintain stable blood sugar, making glucose monitoring a critical component of clinical care.

Keep reading for free.
Create an account to finish this reading and make it part of your study routine.
Continue reading freeAlready have an account? Log in
Updated