


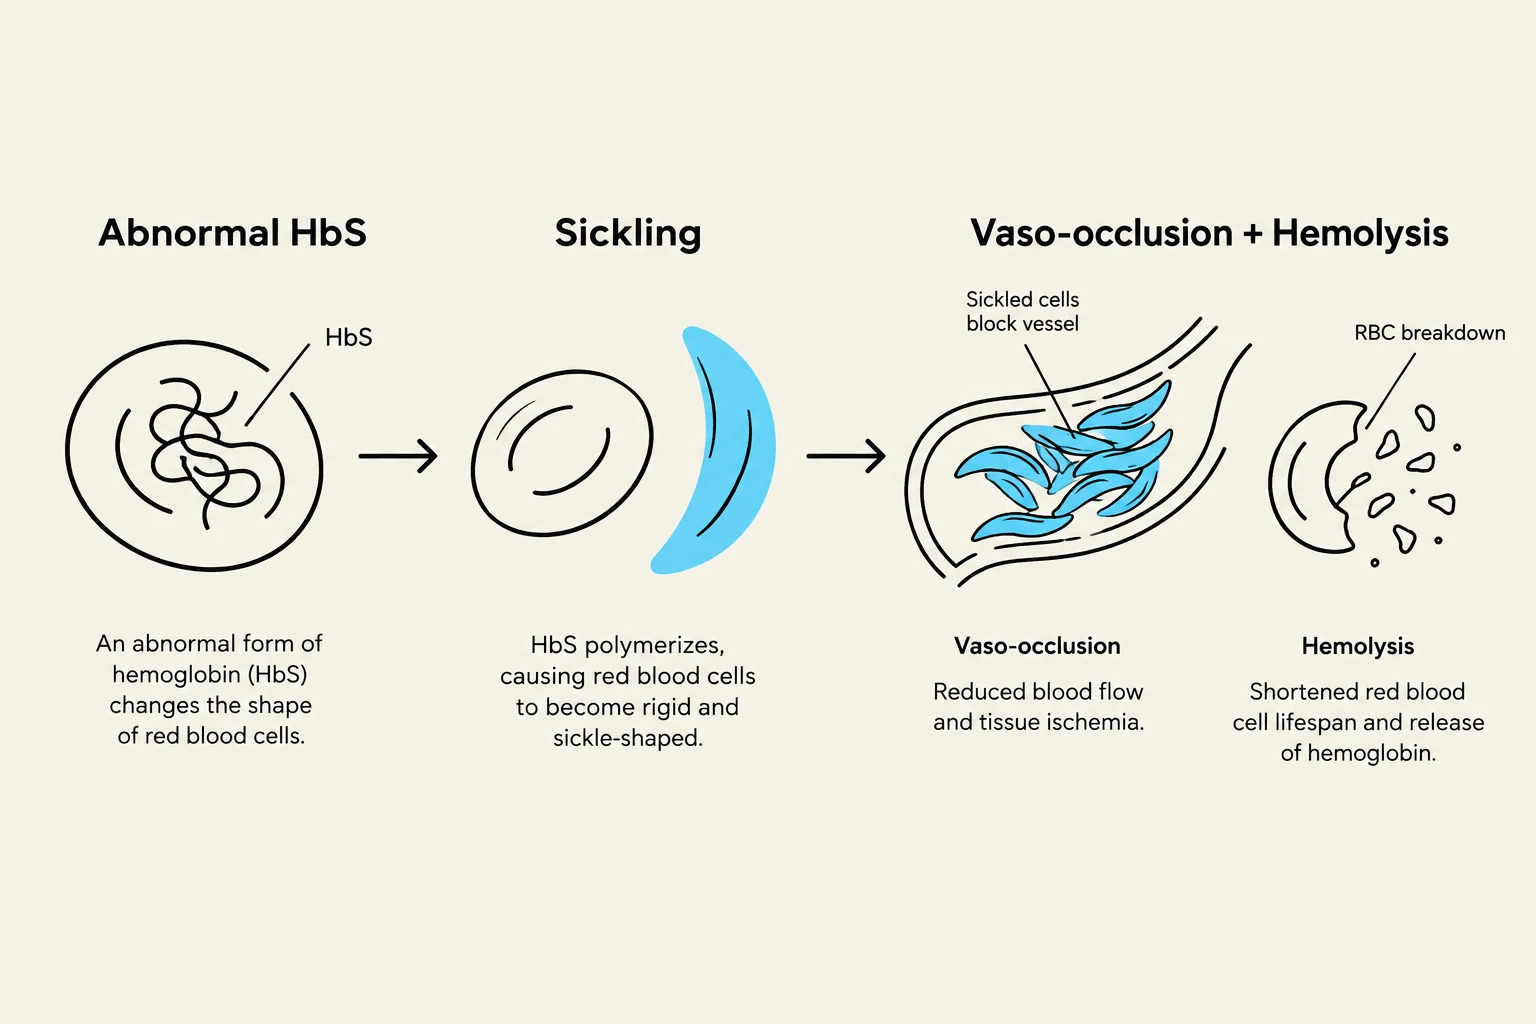
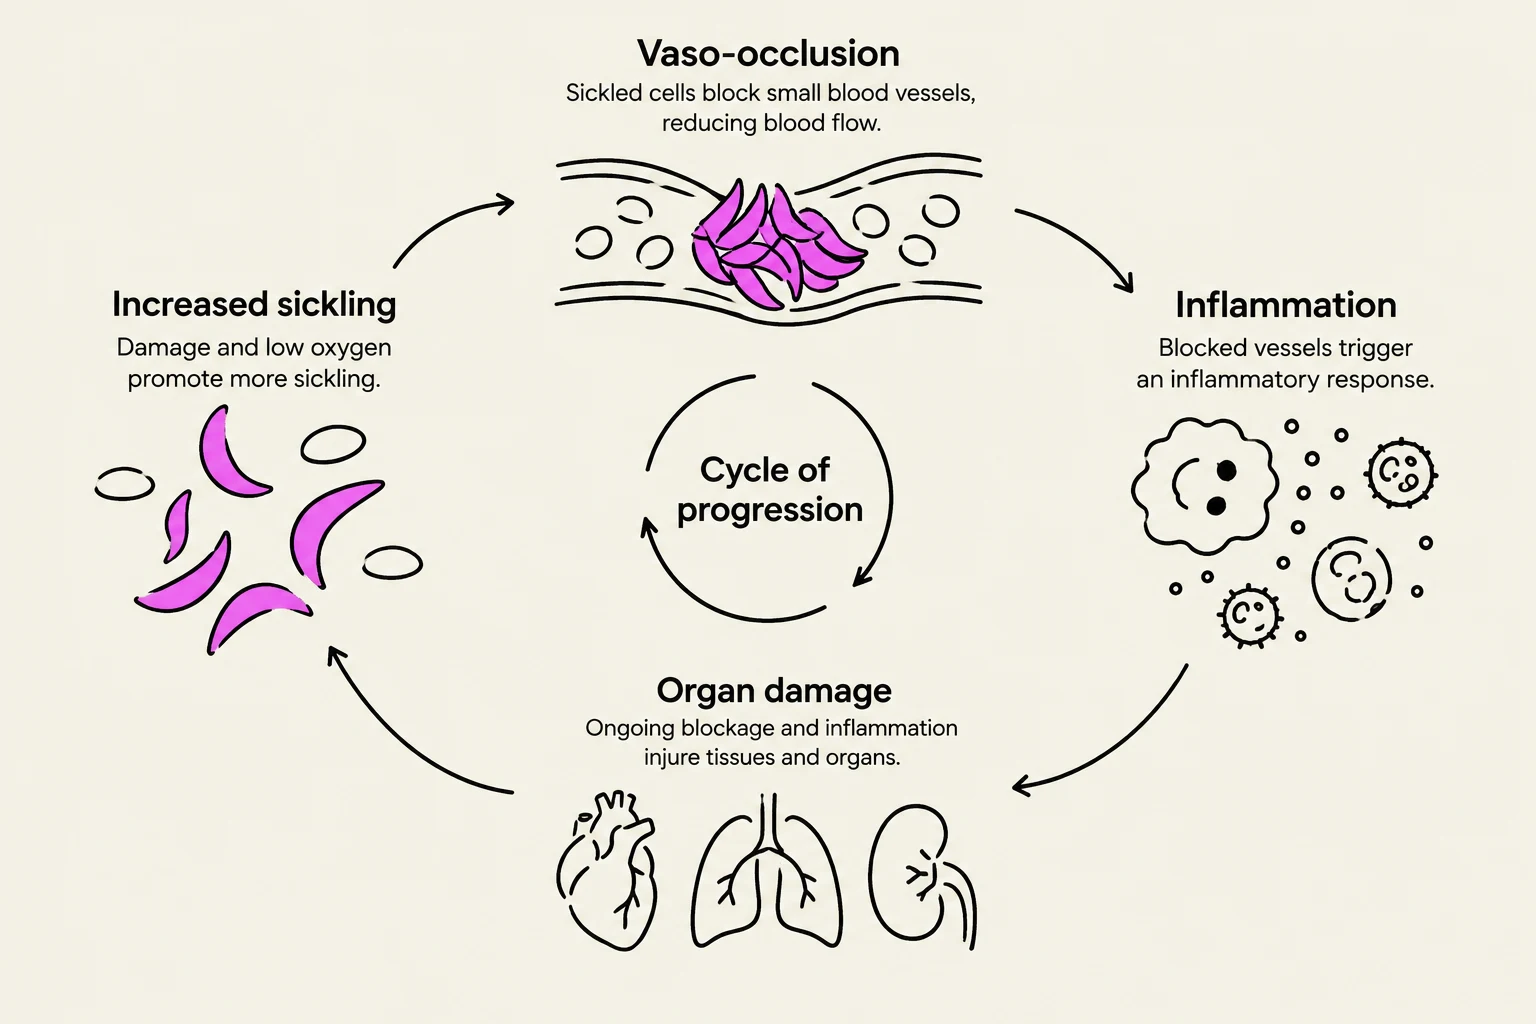
Sickle cell disease (SCD) is a complex, multi-systemic genetic disorder characterized by the production of abnormal hemoglobin. This single molecular defect triggers a cascade of physiological events, ranging from cellular deformation to life-threatening organ damage. Understanding the progression from a genetic mutation to clinical crises is essential for grasping the management of this condition.
Genetics and Etiology
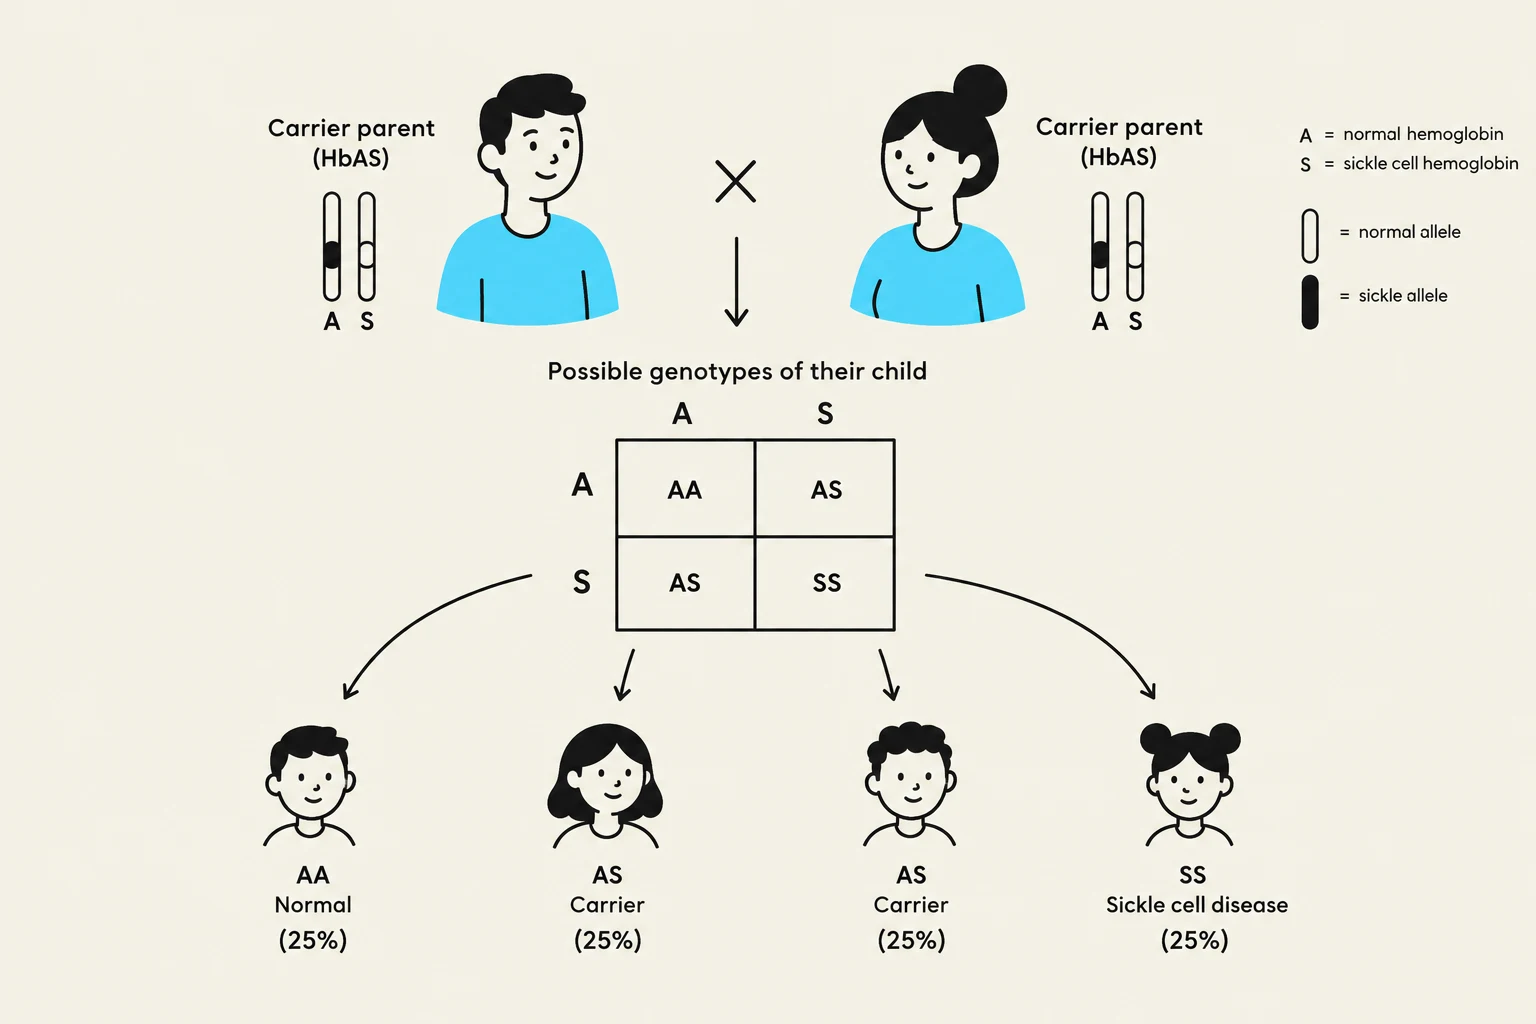
At its core, sickle cell disease is a genetic disorder caused by a specific mutation in the HBB gene, which provides instructions for making a protein called beta-globin. This mutation results in the production of Abnormal HbS (hemoglobin S) instead of the normal adult hemoglobin (HbA).
The inheritance pattern of SCD is autosomal recessive. This means that for an individual to manifest the disease, they must inherit two abnormal HbS genes—one from each parent. Individuals who inherit only one mutated gene are known as carriers (sickle cell trait) and typically do not show significant symptoms, though they can pass the gene to their offspring.
Incidence and Prevalence
The prevalence of the HBB mutation varies significantly across different populations:
- United States: Approximately 100,000 people live with the disease.
- Black/African American populations: Roughly 1 in 13 (8%) births.
- Hispanic-American populations: Approximately 1 in 16,300 births.
- Global Impact: Millions of people are affected worldwide.
Keep reading for free.
Create an account to finish this reading and make it part of your study routine.
Continue reading freeAlready have an account? Log in
Updated