An overview of pharmacokinetics, including absorption, distribution, metabolism, and excretion (ADME) processes, and their impact on drug efficacy and safety. This covers routes of administration, bioavailability, volume of distribution, first-pass metabolism, renal excretion, clearance, and half-life.
In the study of pharmacology, we often distinguish between what a drug does to the body (pharmacodynamics) and what the body does to the drug (pharmacokinetics). While pharmacodynamics focuses on the biochemical and physiological effects of drugs and their mechanisms of action, pharmacokinetics (PK) uses mathematical models to describe the movement of a drug through the body.
To understand how a drug reaches its target and how long it stays there, we use the ADME framework: Absorption, Distribution, Metabolism, and Excretion. Mastering these processes is essential for clinicians to ensure drug efficacy—keeping concentrations above the Minimum Effective Concentration (MEC)—and safety—keeping them below the Maximum Safe Concentration (MSC).
Absorption
Absorption is the movement of an unchanged drug from its site of administration into the systemic circulation. The rate and extent of this process are influenced by the route of administration, the drug's physical properties, and the physiological state of the patient.
Routes of Administration
The route chosen significantly dictates the onset and intensity of the drug's effect:
Parenteral Routes: These include Intravenous (IV), Subcutaneous (SC), and Intramuscular (IM) injections. IV administration provides immediate onset because the drug is injected directly into the circulation, bypassing the absorption phase entirely. SC and IM injections require the drug to diffuse from the injection site into the blood, resulting in a delayed onset but often a more prolonged action.
Oral Dosage Forms: These include tablets, capsules, and liquids. Oral administration is common but subject to the complexities of the gastrointestinal (GI) tract.
The GI Tract and Oral Absorption
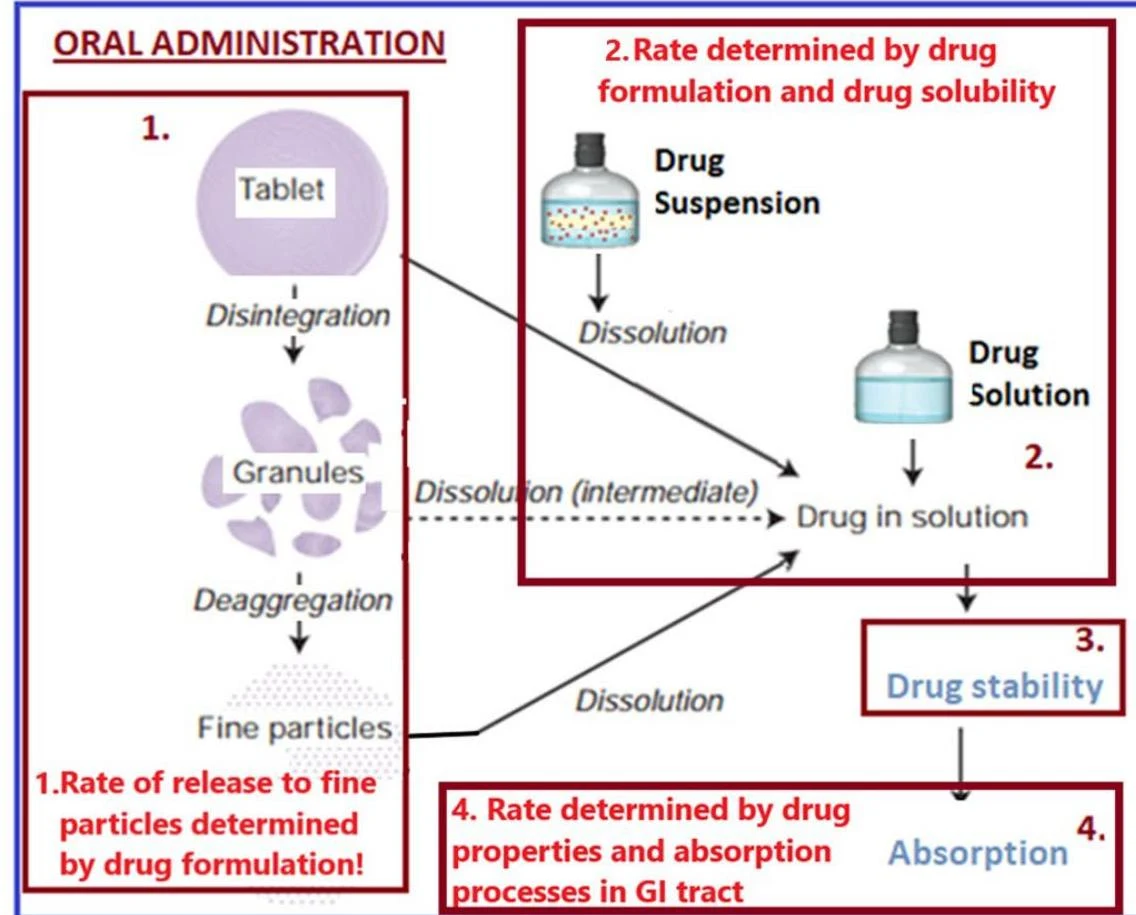
For oral drugs, the process begins with drug dissolution—the breaking down of the dosage form and the dissolving of the drug into the GI fluids.
This diagram illustrates the process of oral drug administration, detailing steps from tablet disintegration to drug absorption in the GI tract, and factors influencing these processes such as drug formulation, solubility, and stability.
The stomach primarily aids in disintegration and dissolution; however, its low pH (1–3) can destroy acid-unstable drugs or enhance the solubility of basic drugs. The small intestine is the primary site for absorption due to its massive surface area (villi and microvilli), moderate pH (6–7.5), and high vascularity.
Mechanisms of Transport
Drugs cross biological membranes through several mechanisms:
Passive Transmembrane Diffusion: The most common method. It is favored by drugs with low molecular weight (< 500) and moderate lipophilicity (Log P between -1 and 5).
Carrier-Mediated Transport: Uses specialized proteins for influx (moving drug into circulation) or efflux (pumping drug back into the GI tract).
Endocytosis: A minor mechanism used for very large or lipophilic molecules.
Exam Tip: Drug Interactions in the Gut
Watch out for complexation and chelation. For example, anion exchange resins like cholestyramine can bind acidic drugs, preventing absorption. Similarly, tetracyclines can form insoluble chelates with polycations like iron (ferrous sulfate) or calcium in dairy, significantly reducing their absorption.
Bioavailability (F)
Bioavailability is the fraction of the administered dose that reaches the systemic circulation in an unchanged form. It is often compared to IV administration, which has a bioavailability of 100% (F=1.0).
The extent of absorption is measured by the Area Under the Curve (AUC), while the rate is characterized by Cmax (peak concentration) and Tmax (time to reach peak).
F=AUCivAUCoral⋅DoseoralDoseiv
Distribution
Once in the blood, the drug undergoes distribution—the process by which it moves from the circulation to various tissues and fluids in the body.
Volume of Distribution (Vd)
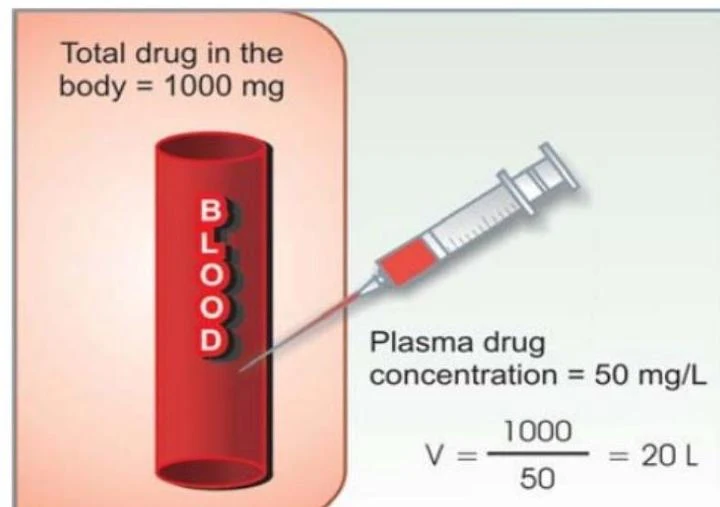
The Volume of Distribution (Vd) is a theoretical volume that relates the total amount of drug in the body to the concentration measured in the plasma.
This illustration demonstrates the calculation of the volume of distribution for a drug in the body. It shows that with a total drug amount of 1000 mg and a plasma drug concentration of 50 mg/L, the volume of distribution is calculated to be 20 L.
A high Vd suggests the drug has distributed extensively into tissues, whereas a low Vd suggests the drug remains primarily in the plasma.
Factors Influencing Distribution
Plasma Protein Binding: Many drugs bind to proteins like albumin. Only the "free" (unbound) drug is pharmacologically active and able to diffuse into tissues. Competition for these binding sites can lead to toxicity.
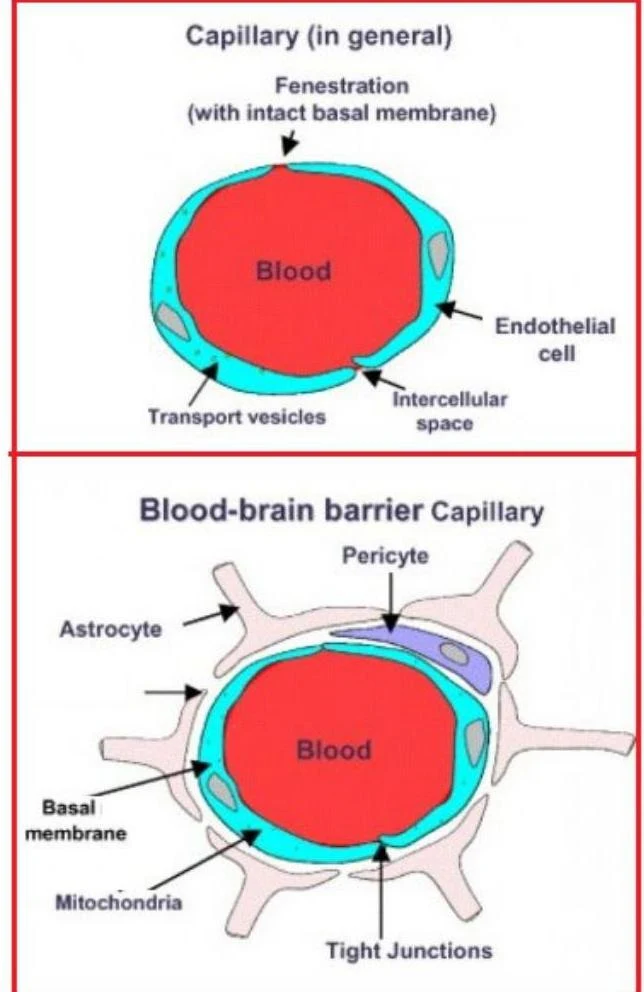
Perfusion and Barriers: Tissues with high blood flow (e.g., brain, heart) receive drugs more rapidly. Furthermore, specialized barriers like the Blood-Brain Barrier (BBB) restrict the movement of many substances.
Diagrams illustrating a general capillary and a blood-brain barrier capillary, highlighting their structural differences and key components like endothelial cells, tight junctions, and astrocytes.
Metabolism
Metabolism is the enzyme-catalyzed chemical transformation of drugs within the body. The liver is the primary organ for this process.
First-Pass Metabolism
A critical concept for oral drugs is first-pass metabolism. When a drug is absorbed from the GI tract, it travels via the portal vein directly to the liver before reaching systemic circulation. If the liver metabolizes a large portion of the drug during this first pass, the oral bioavailability is significantly reduced.
Metabolic Outcomes
Metabolism typically occurs in two phases: Phase I (oxidation, reduction, hydrolysis) and Phase II (conjugation). These processes can result in:
Inactive metabolites: The drug is neutralized and prepared for excretion.
Active metabolites: The drug remains active or becomes more active.
Toxic metabolites: The transformation produces harmful substances (e.g., acetaminophen metabolism).
Clinical Note: Enzyme Induction and Inhibition
Drugs can interact by altering enzyme activity. Inhibitors (like ritonavir or ketoconazole) block enzymes, leading to higher drug levels and potential toxicity. Inducers (like rifampin) increase enzyme production, which can speed up metabolism and cause therapeutic failure.
Excretion
Excretion is the final elimination of drug metabolites from the body. The kidneys are the most important organ for this process.
Renal Excretion Mechanisms
The kidneys utilize three primary processes to remove drugs from the blood:
Glomerular Filtration (GF): Passive filtration of drugs through the glomerulus.
Tubular Secretion (TS): Active transport of drugs from the blood into the renal tubule.
Tubular Reabsorption (TR): The movement of drugs from the tubule back into the blood.
The pH of urine can influence excretion. For example, making urine more alkaline can increase the excretion of acidic drugs like salicylic acid by preventing their reabsorption.
Clearance and Half-life
To quantify how quickly a drug is removed, we use two key parameters:
Clearance (CL): The volume of plasma cleared of the drug per unit of time.
Half-life (t1/2): The time required for the plasma concentration of a drug to decrease by 50%.
t1/2=k0.693
Where k is the elimination rate constant. Understanding the half-life is vital for determining dosing frequency and reaching steady-state—the point where the rate of drug administration equals the rate of elimination.
Summary of Clinical Importance
Pharmacokinetics is not a static science; it varies significantly between patients due to genetics (pharmacogenomics), age, and disease states. For instance, kidney or liver disease can reduce clearance, leading to drug accumulation and toxicity. Conversely, cirrhosis can dramatically increase the bioavailability of certain drugs by reducing first-pass metabolism.
Keep reading for free.
Create an account to finish this reading and make it part of your study routine.